OFFICE NO 5 & 6, 3RD FLOOR,
EARTH ALPHA COMPLEX
VASNA BHAYLI RD, VADODARA
GUJARAT, INDIA

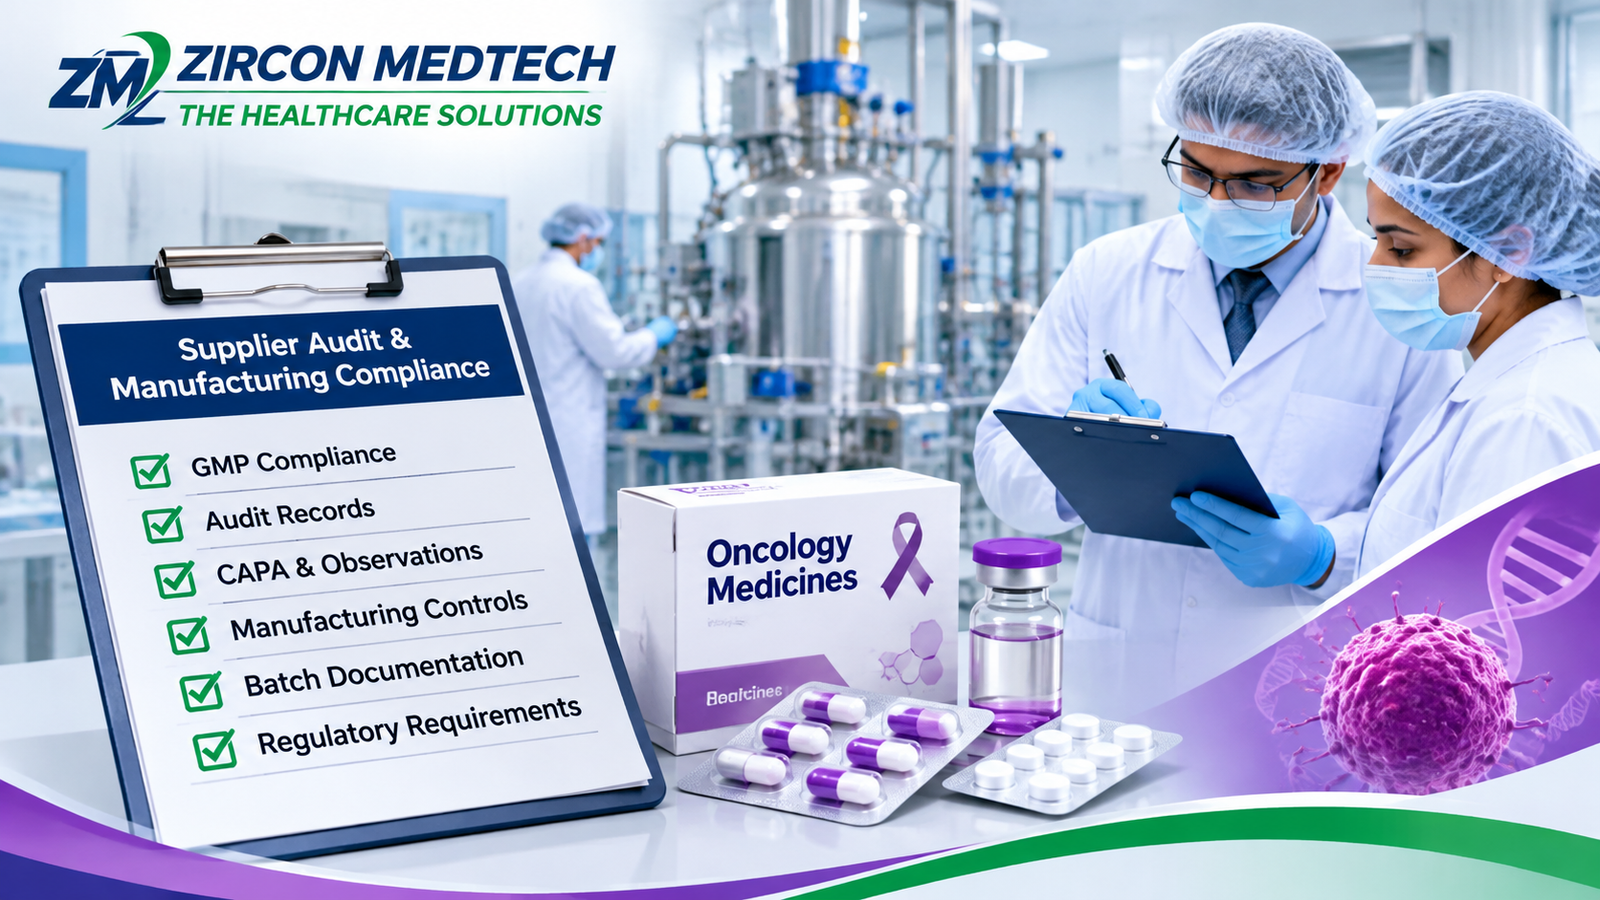
What Should Distributors Check When Selecting a Cancer Medicine Exporter in India?
A cancer medicine exporter in India may supply products manufactured at its own facilities or sourced from approved third-party manufacturers. International buyers should establish which organisation manufactures each product and which party is responsible for export documentation and quality-related communication.
Supplier evaluation should cover the exact manufacturing site and product rather than relying only on general company credentials.
For example, a manufacturer may hold a GMP certificate for one facility, but that certificate may not cover every dosage form, production line or product proposed for export.
Common Reasons to Review Oncology Medicine Suppliers
Why Are Supplier Audit Records Important for Oncology Medicine Exports?
Supplier audit records provide evidence of how a manufacturer or supplier's quality systems have been evaluated.
An audit may examine manufacturing processes, personnel training, documentation practices, equipment qualification, contamination controls and the handling of deviations.
For oncology medicines, the review may also need to consider hazardous material handling, containment measures and product-specific manufacturing risks.
A completed audit does not automatically demonstrate that every concern has been resolved. Distributors should examine the findings, corrective actions and available evidence of closure.
A Supplier Audit Review Should Establish
Verify the Manufacturer and Manufacturing Site Details
Before reviewing audit documentation, confirm the identity of the legal manufacturer and the facility responsible for producing the medicine.
A supplier may work with multiple manufacturers, and different products may come from different sites.
Distributors should request site-specific information for the exact medicine presentation being considered.
Manufacturer Information to Confirm
Where the exporter and manufacturer are separate organisations, clarify which party provides documents, manages complaints and communicates manufacturing changes.
Review Good Manufacturing Practice Compliance
Good Manufacturing Practice is a central consideration when evaluating pharmaceutical manufacturers.
GMP requirements address the systems needed to manufacture and control medicines consistently according to applicable quality standards.
International buyers should assess the standards recognised by the destination market and verify that the evidence applies to the relevant manufacturing site and activities.
The World Health Organization publishes GMP guidance covering pharmaceutical quality systems, production, quality control, documentation and other manufacturing requirements.
GMP Compliance Areas to Evaluate
|
Compliance Area |
What Distributors Should Review |
|
Pharmaceutical quality system |
Quality responsibilities, procedures and management oversight |
|
Manufacturing facilities |
Premises, production areas and appropriate environmental controls |
|
Equipment qualification |
Qualification, calibration and maintenance records |
|
Personnel training |
Training records and role-specific competency |
|
Documentation practices |
Controlled procedures, batch records and data integrity |
|
Quality control |
Testing methods, specifications and laboratory controls |
|
Deviation management |
Investigation procedures and documented outcomes |
|
Change control |
Evaluation and approval of manufacturing changes |
|
Product traceability |
Batch identification and relevant distribution records |
A GMP certificate should be assessed for its issuing authority, scope, validity and relevance to the specific site. Where appropriate, verify the information with the issuing authority.
Examine Supplier Audit Reports and Inspection Findings
Audit reports should provide enough information to understand the scope of the assessment and the manufacturer's response to identified concerns.
Some reports contain confidential manufacturing information. Distributors may receive an audit summary, redacted findings or other appropriate evidence depending on contractual arrangements and quality agreements.
The review should focus on the significance of findings rather than simply counting the number of observations.
Information to Review in Supplier Audit Records
An older audit may need to be supplemented with more recent inspection information, particularly when manufacturing processes, facilities or regulatory status have changed.
Assess Corrective and Preventive Action Records
Corrective and Preventive Action (CAPA) records show how manufacturers investigate and address quality problems.
A manufacturer may identify an issue during an internal audit, regulatory inspection, complaint investigation or routine production review.
The existence of a CAPA plan does not establish that the underlying problem has been resolved.
Distributors should check whether actions have been completed and whether their effectiveness has been evaluated.
Questions to Ask When Reviewing CAPA Records
Unresolved critical findings should be escalated to qualified quality personnel before a supplier or affected product is approved.
Evaluate Oncology-Specific Manufacturing Controls
Cancer medicines may require specialised manufacturing controls depending on the active pharmaceutical ingredient, dosage form and associated hazards.
Some oncology products involve highly potent or cytotoxic substances that require particular attention to containment, occupational safety and cross-contamination prevention.
The necessary controls should be based on a documented product- and process-specific risk assessment.
Manufacturing Controls to Review
For sterile oncology injectables, the assessment should additionally address applicable sterile manufacturing requirements, including contamination control, aseptic processing or sterilisation, and relevant validation activities.
Not all oncology medicines require the same controls. The review must reflect the particular formulation and manufacturing process.
Check Product-Specific Quality Documentation
Supplier-level compliance evidence should be supported by appropriate documentation for each medicine being sourced.
A manufacturer may have an established quality system, but the distributor still needs to verify that the proposed product meets the applicable requirements of the destination market.
Documents to Request Where Applicable
|
Document |
Purpose of Review |
|
Manufacturing licence |
Confirm relevant manufacturing authorisation |
|
GMP certificate or inspection evidence |
Review site-specific GMP compliance |
|
Product authorisation documents |
Confirm the applicable product approval status |
|
Certificate of Analysis |
Review batch-specific test results against specifications |
|
Product specifications |
Identify approved quality requirements |
|
Stability information |
Support approved shelf life and storage conditions |
|
Batch documentation |
Support batch identity and manufacturing traceability |
|
Packaging and labelling information |
Confirm the proposed market presentation |
|
Quality agreement |
Define responsibilities between relevant parties |
Document requirements vary by country, product and transaction. The local importer and regulatory team should determine which records are necessary before ordering, dispatch and market release.
Review Batch Records and Quality Control Arrangements
Batch documentation helps establish how a particular quantity of medicine was manufactured, tested and evaluated.
Distributors may not routinely receive complete manufacturing batch records. However, the responsible quality team should establish what batch-specific evidence is required and how additional records can be accessed when necessary.
Batch-Related Information to Verify
For oncology injectables, additional product-specific evidence may be required concerning sterility assurance, particulate contamination and other relevant quality attributes.
Assess Data Integrity and Documentation Practices
Reliable manufacturing records are essential for demonstrating that pharmaceutical production and testing activities were performed appropriately.
Distributors should consider whether the manufacturer's documentation systems support accurate, complete and traceable records.
Data integrity expectations commonly include the ALCOA principles and their extended form, ALCOA+, which address characteristics such as attributable, legible, contemporaneous, original, accurate, complete, consistent, enduring and available data.
Documentation Controls to Examine
Repeated unexplained documentation discrepancies should trigger further review rather than being treated as routine administrative errors.
Confirm Destination Market Regulatory Requirements
A cancer medicine exporter in India may supply products to several international markets, but regulatory requirements are not necessarily identical across countries.
Before sourcing, the distributor should work with its local regulatory team to confirm the requirements applicable to the product, manufacturer and destination market.
A medicine manufactured and authorised in India is not automatically approved for sale in another country.
Information to Review With the Local Importer
For example, a Brazilian importer should confirm the applicable requirements of ANVISA before relying on a proposed Indian manufacturer or product presentation.
Establish a Supplier Qualification and Approval Process
A documented supplier qualification process helps distributors evaluate manufacturing compliance consistently.
The process should define the information required, the people responsible for reviewing it and the circumstances under which a supplier can be approved.
Suggested Supplier Qualification Process
The process should allow the quality team to reject, suspend or place conditions on a supplier when evidence is insufficient.
Develop a Cancer Medicine Supplier Compliance Checklist
A supplier compliance checklist helps procurement, quality assurance and regulatory teams review the same information before approving a sourcing arrangement.
Suggested Supplier Assessment Checklist
|
Review Area |
Evidence to Confirm |
|
Manufacturer identity |
Legal name, site address and manufacturing responsibilities |
|
Manufacturing authorisation |
Relevant licences and permitted activities |
|
GMP compliance |
Applicable certificates, inspection status and scope |
|
Supplier audit |
Audit date, scope, findings and report status |
|
CAPA review |
Completion and effectiveness of significant actions |
|
Oncology manufacturing controls |
Appropriate containment and contamination prevention |
|
Product quality |
Specifications, testing arrangements and stability evidence |
|
Batch traceability |
Batch identification and relevant quality records |
|
Regulatory suitability |
Destination-market requirements and approvals |
|
Change management |
Notification and approval arrangements |
|
Quality agreement |
Defined responsibilities and escalation procedures |
Keep the completed assessment with supporting records and review it when significant changes occur.
Hypothetical Example of Reviewing an Indian Oncology Medicine Supplier
An international pharmaceutical distributor is evaluating an Indian exporter for two oncology medicine presentations.
The exporter provides manufacturing details, GMP documentation and a summary of a recent supplier audit.
During the review, the distributor identifies an outstanding corrective action concerning equipment cleaning documentation. It also discovers that the GMP evidence supplied does not clearly identify whether the proposed injectable production line is within scope.
The distributor refers both matters to its quality team before confirming supplier approval.
How the Distributor Reviews the Findings
|
Issue |
Review Action |
|
Manufacturing site details |
Confirm the actual manufacturer and relevant production areas |
|
GMP scope uncertainty |
Request evidence covering the proposed manufacturing activities |
|
Outstanding audit observation |
Review the finding, potential product impact and CAPA status |
|
Cleaning documentation concern |
Obtain appropriate corrective action and effectiveness evidence |
|
Injectable product requirements |
Confirm relevant sterile manufacturing compliance |
|
Supplier approval |
Document the quality decision after reviewing the outstanding evidence |
This example demonstrates why supplier approval should be based on the relevance and completeness of quality evidence rather than the availability of a certificate alone.
Common Mistakes to Avoid When Assessing Cancer Medicine Exporters
Relying Only on a GMP Certificate
Verify the issuing authority, manufacturing site, certificate scope and relevance to the proposed products.
Ignoring Unresolved Supplier Audit Findings
Review the significance of observations and obtain appropriate evidence of corrective action before approval.
Assuming Every Product Comes From the Same Manufacturing Facility
Confirm the actual manufacturing site for each product and dosage form.
Overlooking Oncology-Specific Manufacturing Risks
Assess containment, contamination prevention and sterile manufacturing requirements where applicable.
Accepting Incomplete Product Documentation
Confirm the necessary specifications, batch evidence and regulatory documents before the relevant supply stage.
Failing to Monitor Manufacturing Changes
Establish procedures for communicating and evaluating changes to sites, processes, testing arrangements or product presentations.
Discuss Your Oncology Medicine Sourcing Requirements With Zircon Medtech
International distributors looking for a cancer medicine exporter in India should begin with a clear product and supplier assessment plan.
When contacting Zircon Medtech, provide the destination country, required oncology medicines, strengths, dosage forms, expected volumes and preferred delivery schedule.
Include the manufacturing and quality documents required by your organisation, along with applicable regulatory, packaging, storage and shelf-life requirements.
This information can support discussions about available sourcing options and the evidence needed for your supplier qualification process.
Visit Zircon Medtech
Keeping Your Oncology Supplier Compliance Review Focused
Selecting a cancer medicine exporter in India requires a structured evaluation of manufacturing compliance, supplier audit findings and product-specific quality evidence.
Begin by confirming the actual manufacturer and site. Review applicable GMP records, significant audit observations, corrective actions and the controls relevant to the proposed oncology products.
Work with qualified quality and regulatory personnel to resolve outstanding issues and confirm destination-market requirements before approving a supplier or shipment.
A documented supplier qualification process can help international pharmaceutical distributors make informed sourcing decisions while maintaining appropriate expectations for medicine quality, traceability and regulatory compliance.






Share This News